Lysosomal Membrane Permeability On Parkinson's Disease
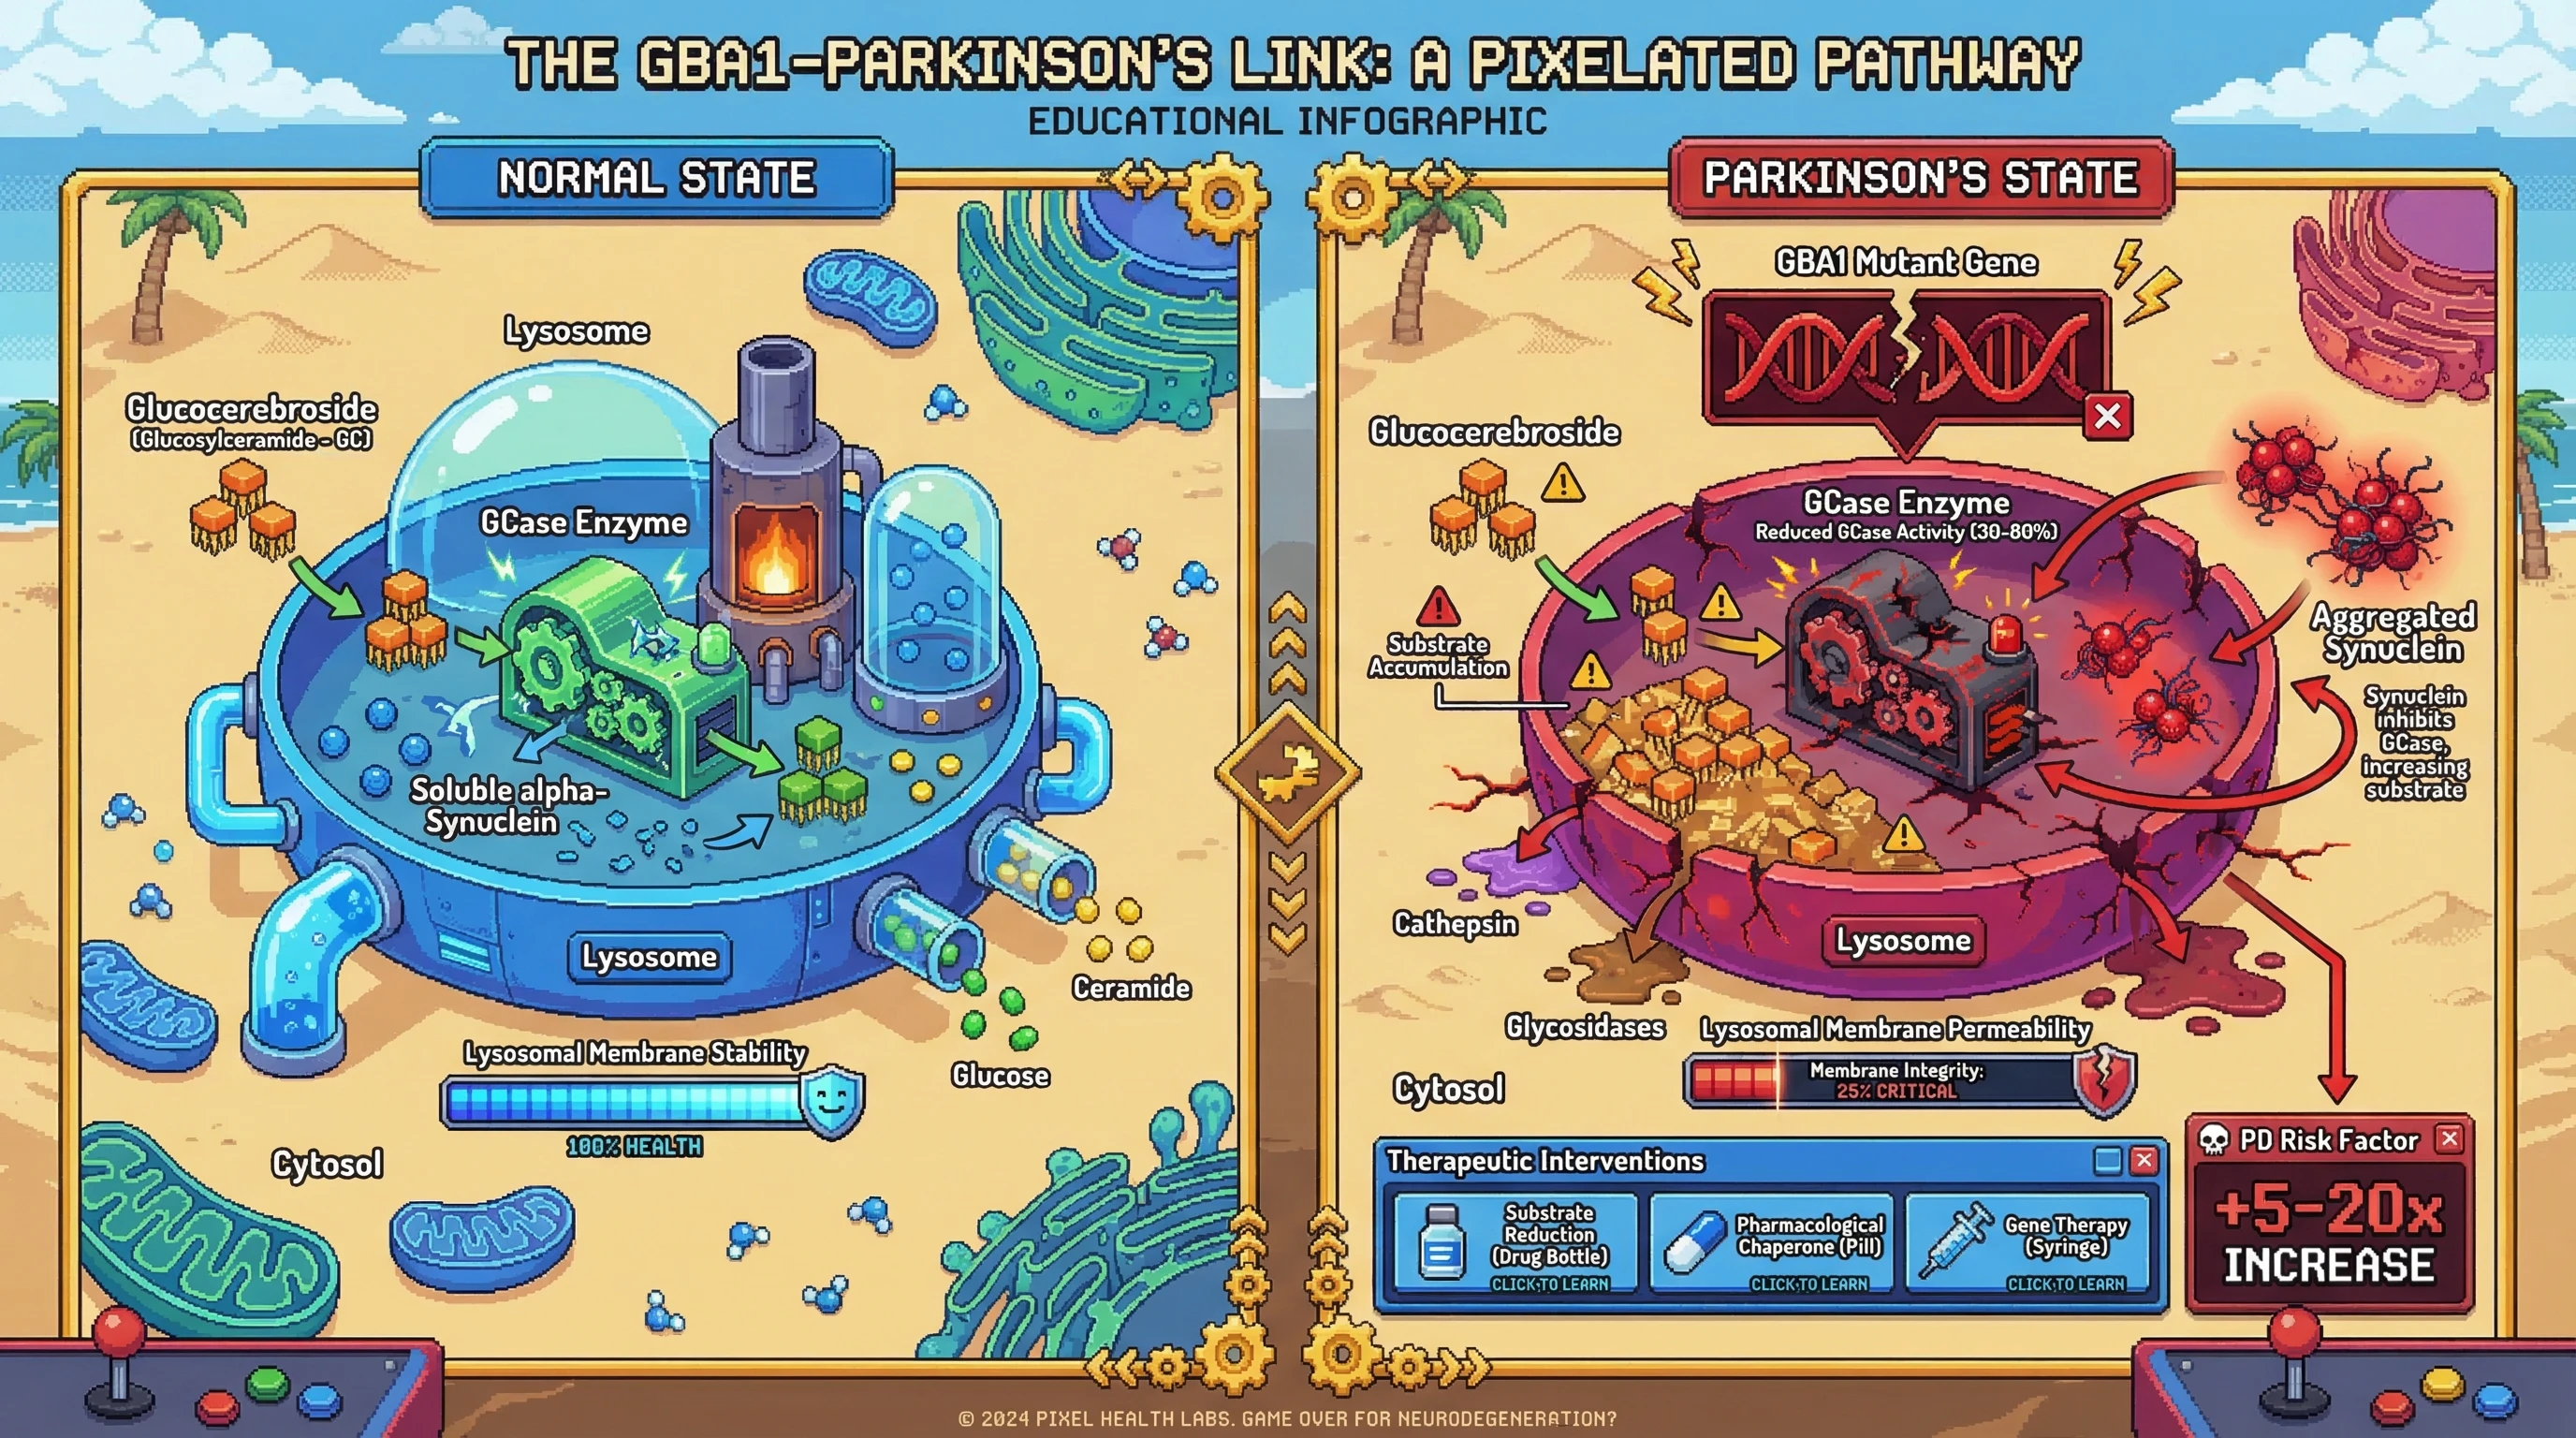
The Central Proposition: GBA1 mutations represent the strongest genetic risk factor for Parkinson's. This hypothesis explains why lipid metabolism failure turns lysosomes from cellular recyclers into bomb fragments.
The Mechanism:
Enzymatic Failure: GBA1 encodes glucocerebrosidase (GCase), which cleaves glucocerebroside into glucose and ceramide inside lysosomes. Mutations reduce GCase activity by 30-80%, depending on allele.
Substrate Accumulation: Uncleaved glucocerebroside accumulates within lysosomal lumen. This glycosphingolipid has biophysical properties that intercalate into membranes.
Membrane Integration: Excess glucocerebroside incorporates into the lysosomal lipid bilayer, altering its biophysical properties increasing rigidity, reducing stability, and changing lipid raft organization.
Permeabilization: Altered membrane becomes susceptible to rupture. Lysosomal membrane permeability increases, leaking contents (cathepsins, glycosidases) into cytosol.
Synuclein Connection: Acidic luminal pH normally keeps alpha-synuclein soluble. Leaked synuclein encounters neutral cytosolic pH, promoting aggregation. Additionally, glucocerebroside directly binds synuclein, stabilizing oligomeric intermediates.
Bidirectional Amplification: Aggregated synuclein inhibits GCase trafficking to lysosomes, further reducing enzymatic activity creating feed-forward loop. Membrane permeability also releases calcium, triggering mitochondrial damage.
The Genotype-Phenotype:
Homozygous GBA1 mutations cause Gaucher's (visceral)
Heterozygous mutations increase PD risk 5-20x
Carrier frequency 5-10% in PD populations
Therapeutic Implications:
Substrate reduction therapy decreasing glucocerebroside synthesis
Pharmacological chaperones stabilizing mutant GCase
Gene therapy delivering functional GBA1 to CNS
Lysosomal membrane stabilizers preventing rupture
This reframes GBA-PD as lysosomal storage disorder of neurons lipids driving synuclein pathology.